New Product Introduction (NPI)
G&H accelerates new product introduction (NPI) for medical devices and life-science instruments - bridging design and manufacturing through validation, risk management, and process excellence.
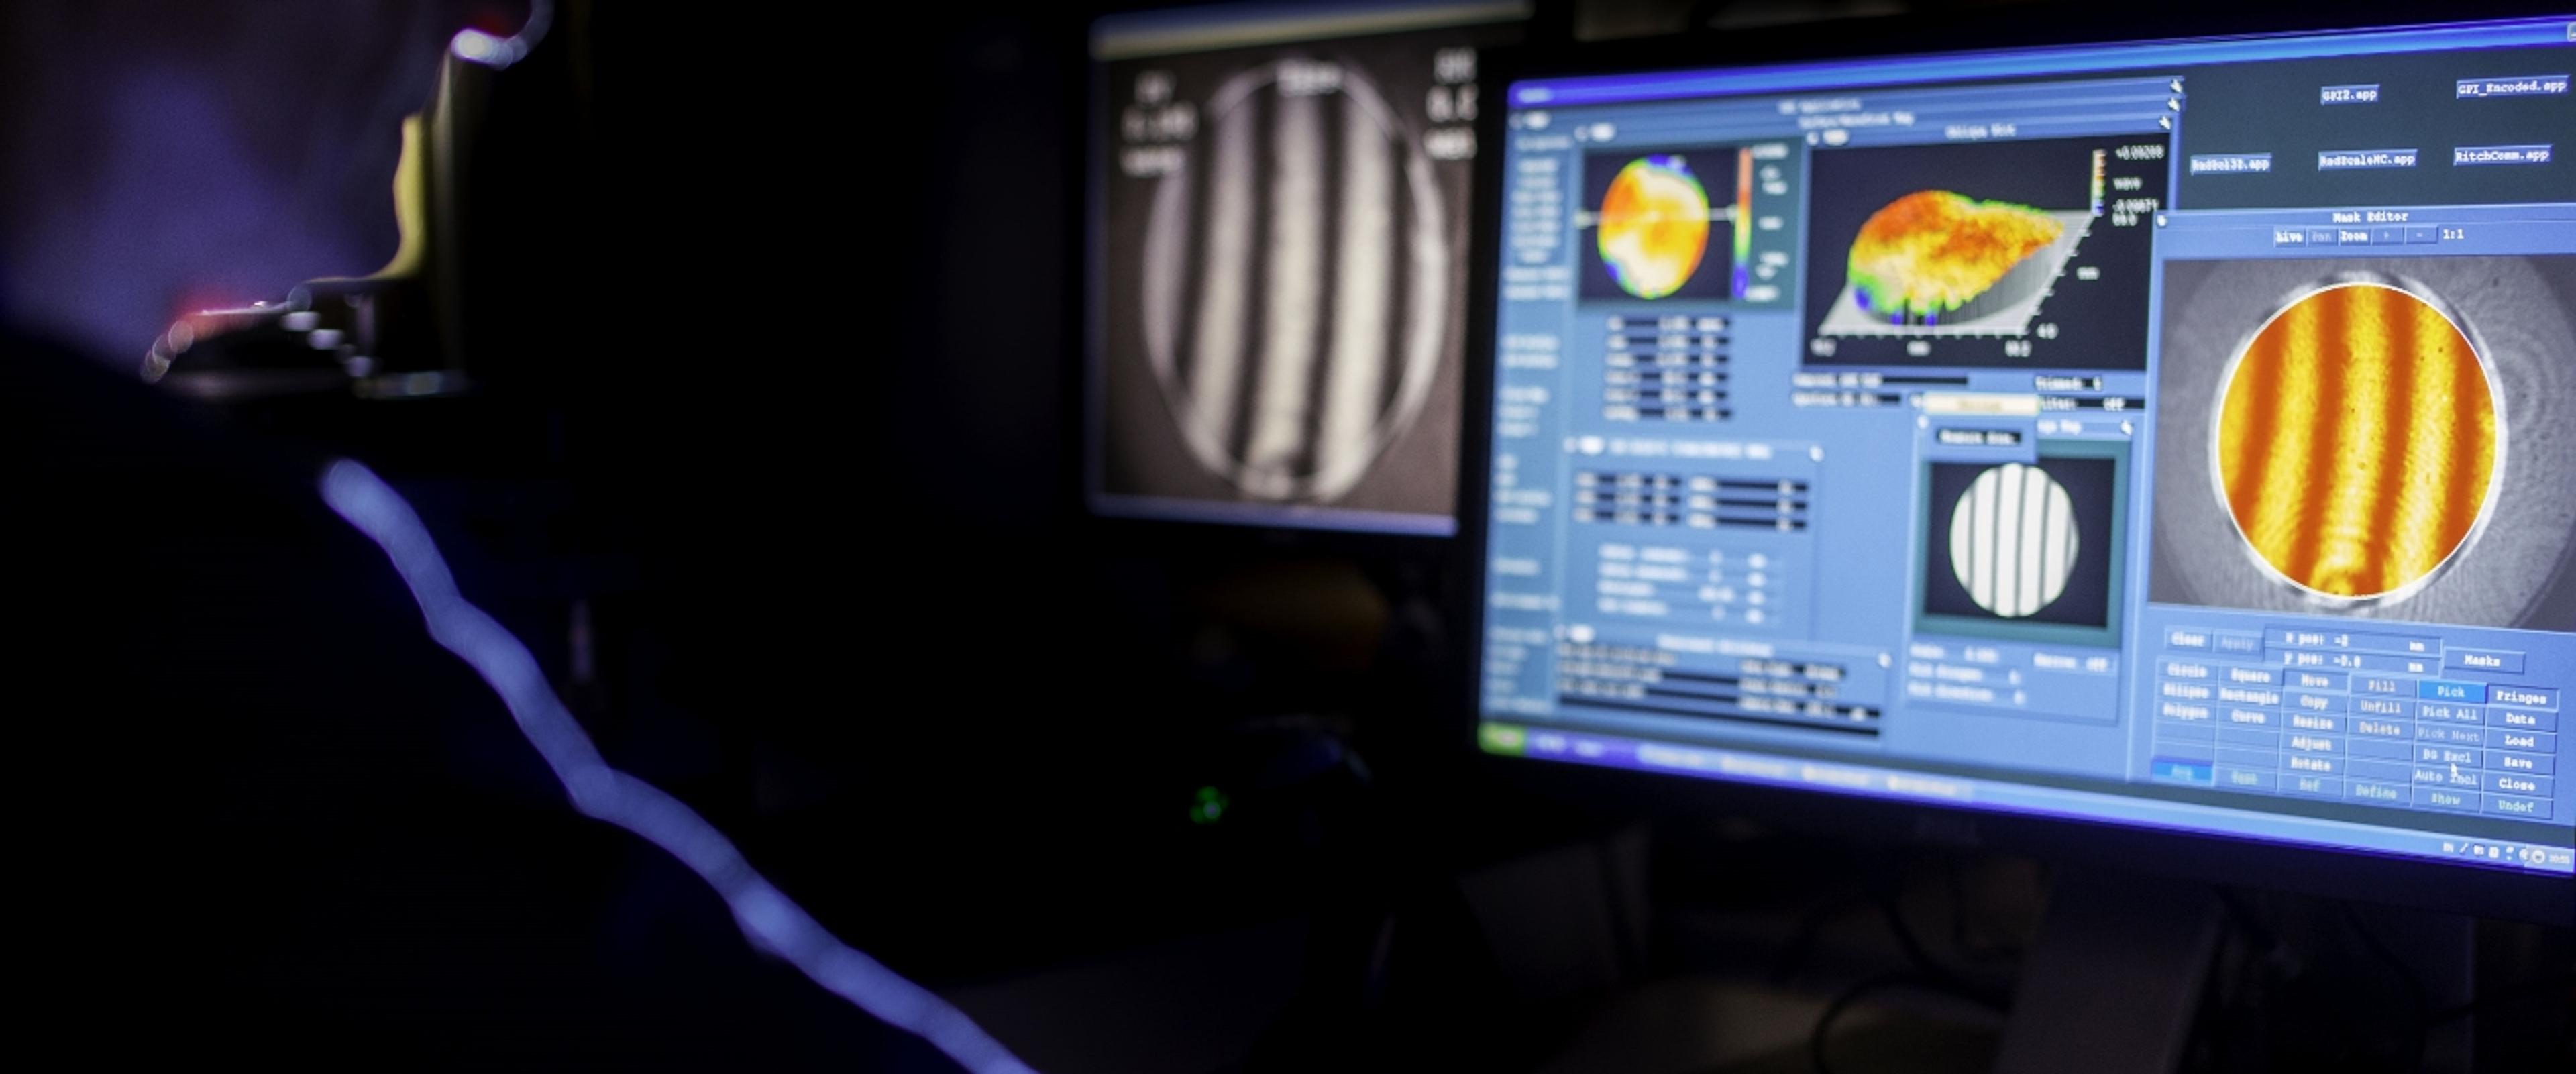

The Validation Manufacture phase is the final step in G&H’s New Product Introduction (NPI) process. This critical stage verifies that every aspect of production - from materials and operators to equipment and documentation - consistently produces devices that meet specification and regulatory requirements.
By completing this validation under ISO 13485 and FDA 21 CFR 820, G&H confirms that the manufacturing process is fully reproducible, controlled, and ready for commercial release.
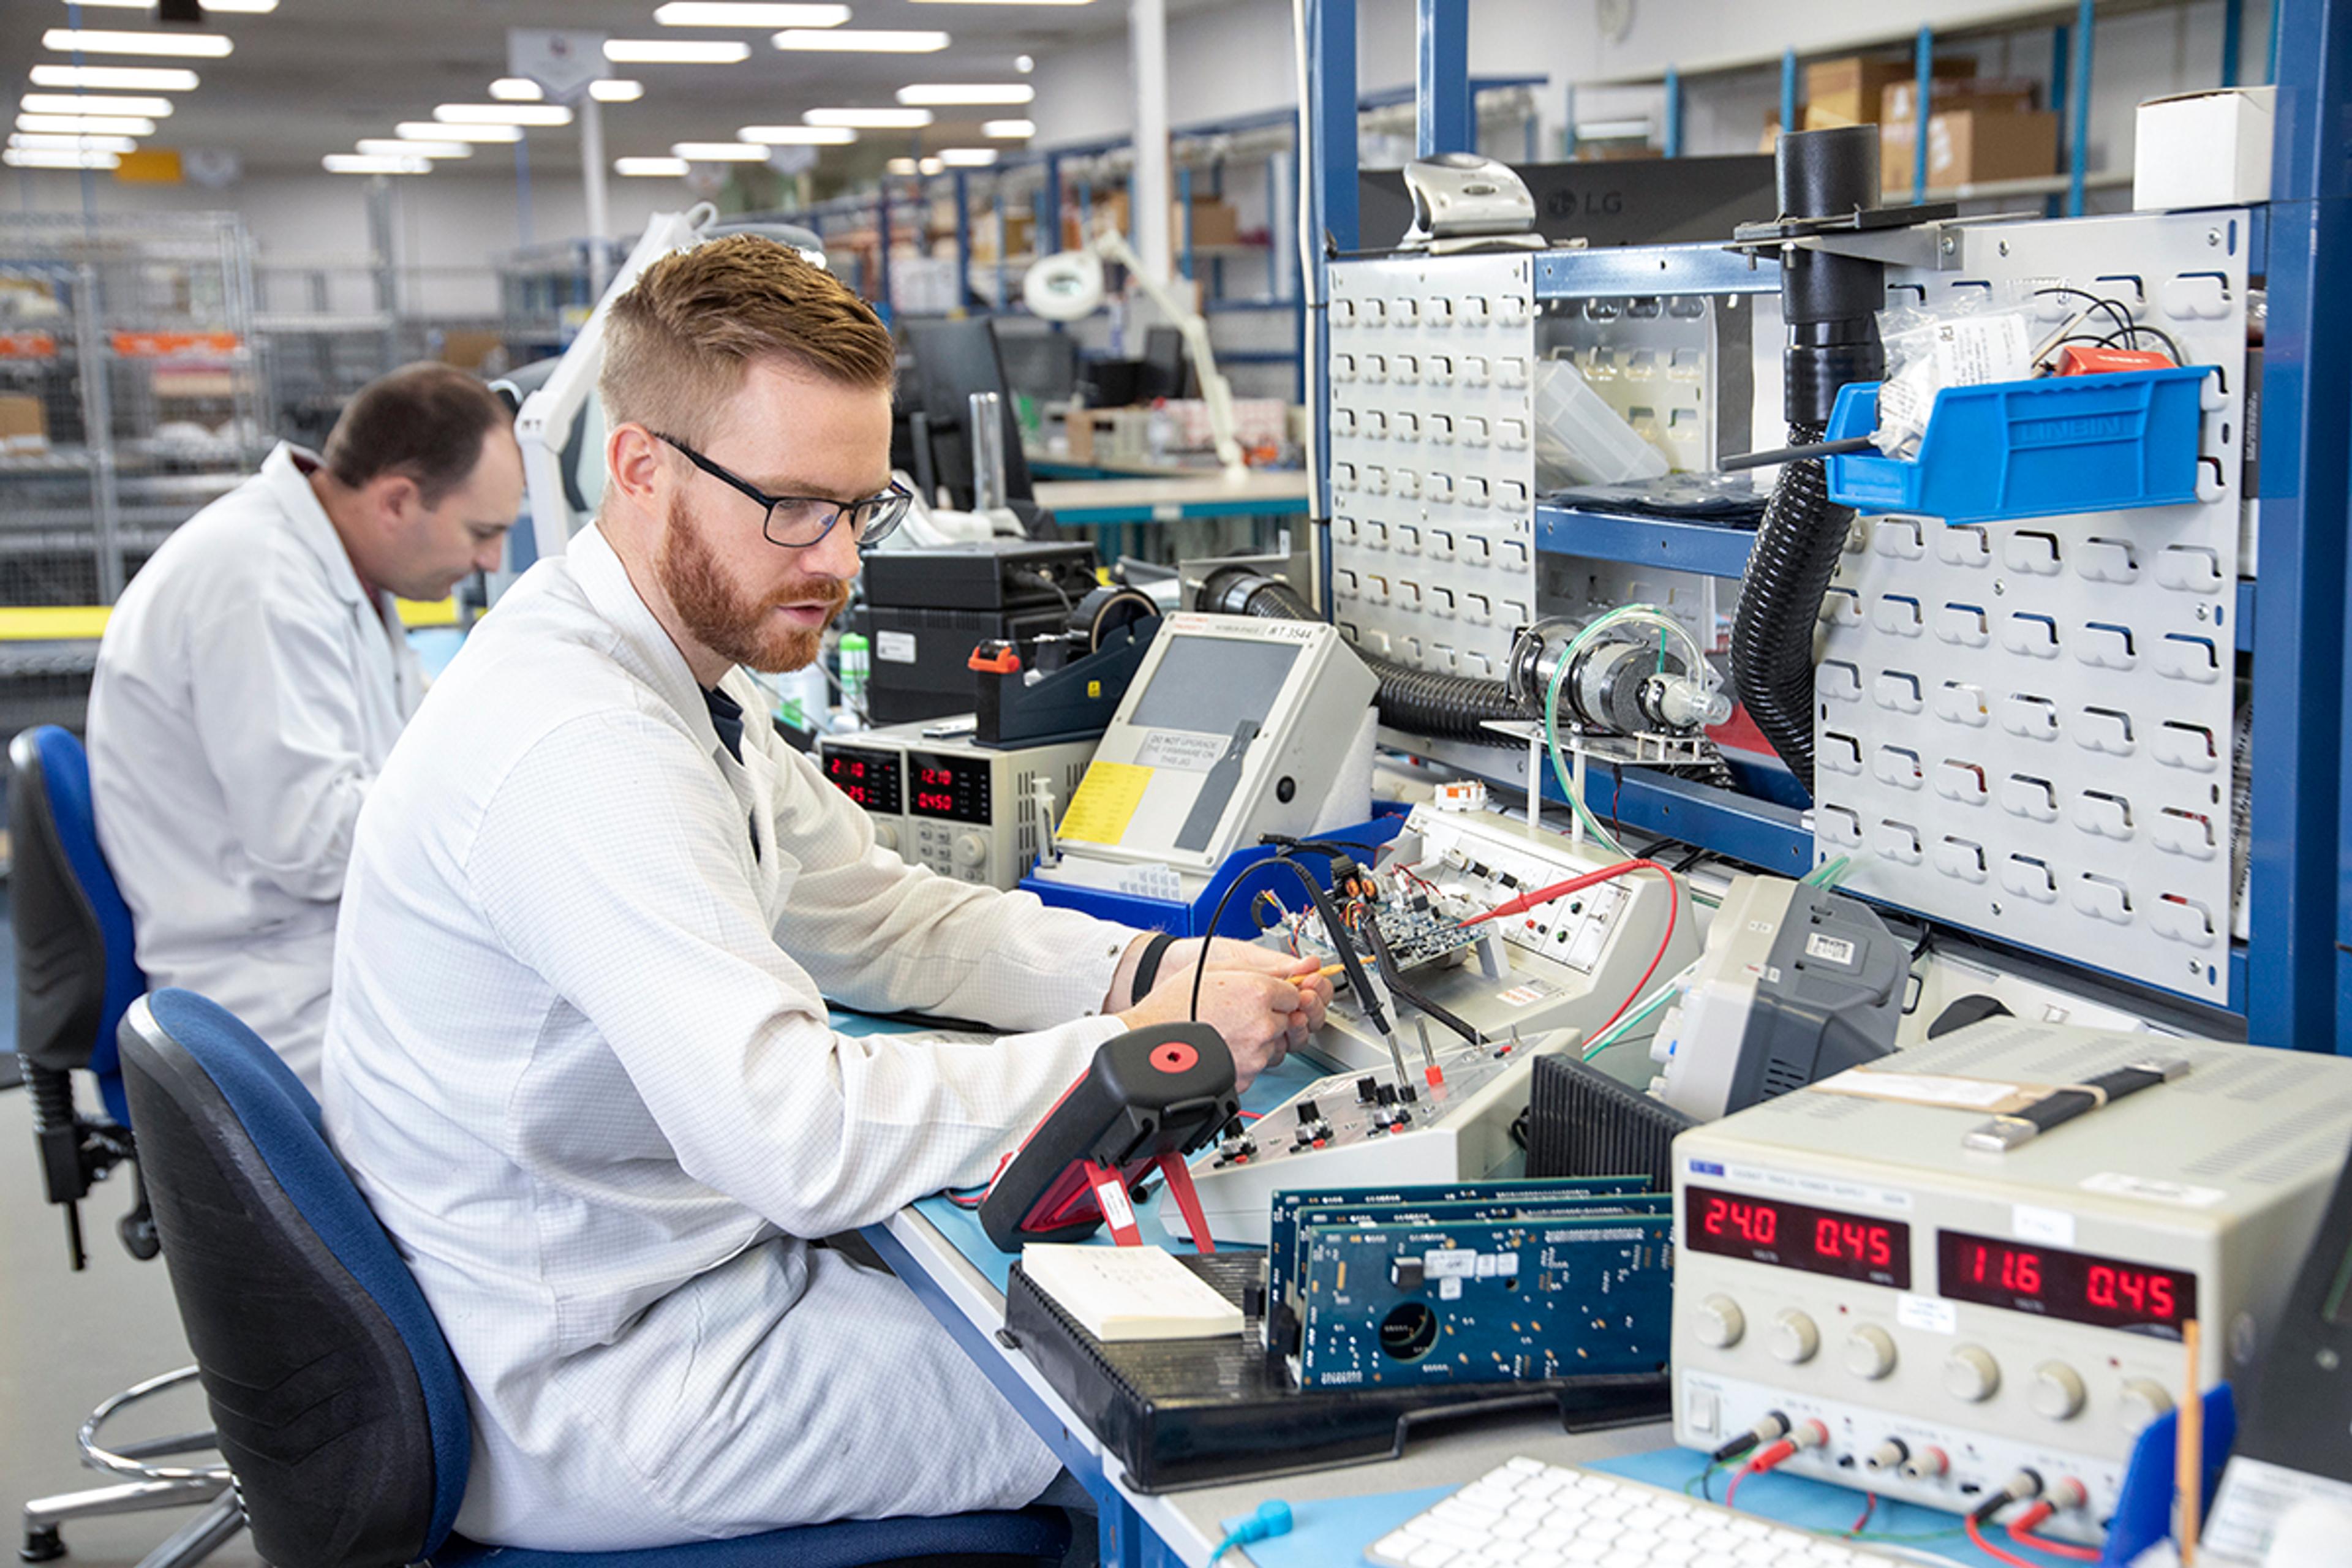
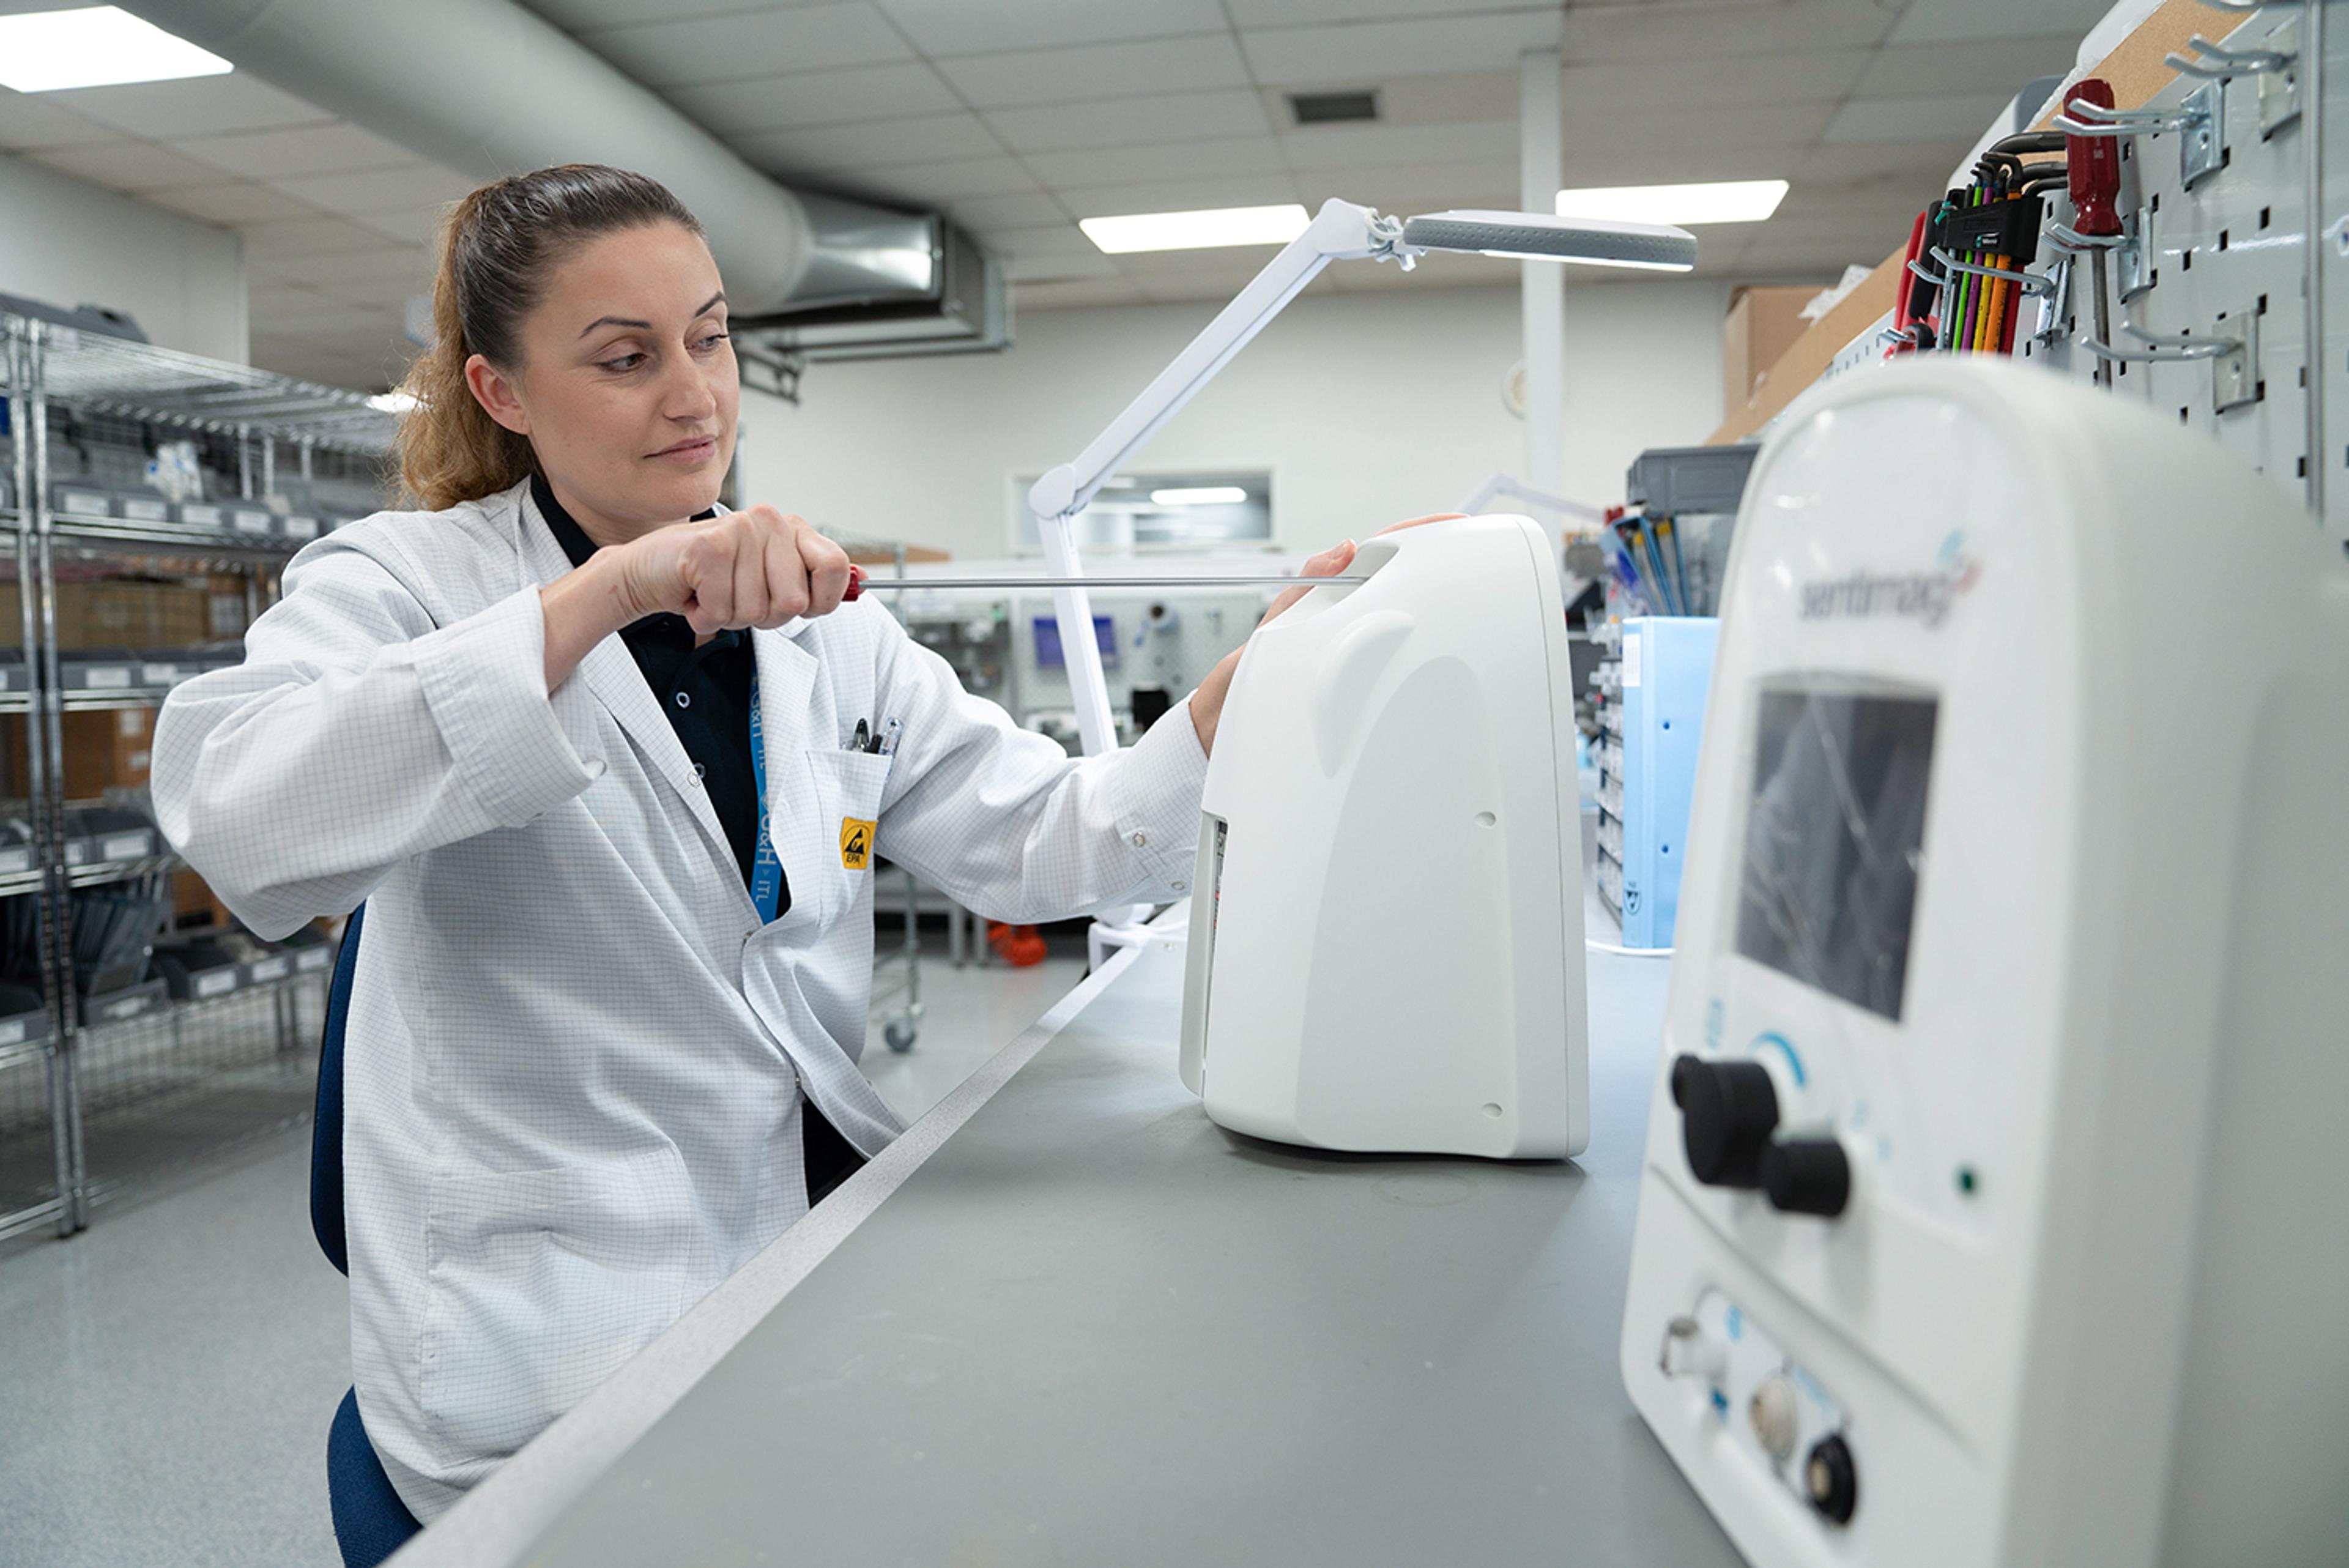
Validation builds are carried out by trained Production Operators under defined conditions using approved tools, materials, and validated instructions. Unlike previous stages, Production Engineering and Subject Matter Experts may observe the process but cannot directly assist with builds, ensuring that results reflect true production capability.
Sampling rationale follows ISO 13485:2016 and statistical best practice - typically three independent test data points or a minimum of 30 devices - to demonstrate process performance and reproducibility.
All results are recorded, reviewed, and approved within G&H’s Quality Management System, ensuring audit readiness and long-term compliance.
Through this structured validation process, G&H | ITL ensures that your medical device is not only manufacturable - but manufactured right, every time.